A contamination finding needs a quality assessment. Product disposition, investigation, cleaning and testing may all need review. Assess the scope using site records rather than a generic cost estimate.
Beyond the lost batch
Suppose monitoring identifies contamination in a manufacturing area. The quality team needs to assess affected equipment, materials and activities. Review the production schedule and any holds under the approved procedures.
For patient-specific products, include clinical supply responsibilities in the impact assessment. Let the responsible quality and clinical teams determine whether notification or other action is needed.
Investigation and staff time
Follow the site investigation procedure. Assign responsibilities across quality, operations, facilities and microbiology as appropriate. Record evidence, conclusions and any corrective actions before closing the investigation.
When estimating investigation resources, include record review, sampling, testing and specialist assessment. Define which activities are needed for the event rather than assuming a standard duration.
Regulatory exposure and schedule slippage
Review individual findings together with monitoring trends. Under EU GMP Annex 1, monitoring contributes to assessment of contamination control. The quality team should evaluate repeated findings and determine investigation and escalation requirements.
Assess potential schedule effects with the teams responsible for manufacturing and regulatory obligations. Use the actual process and investigation evidence. A contamination finding alone does not establish a clinical hold or a particular delay.
Quality records in partnership reviews
Include quality records in supplier and partnership reviews where relevant. Ask how findings are investigated, how corrective actions are checked and how records are shared under the agreed confidentiality terms.
Review the prevention program
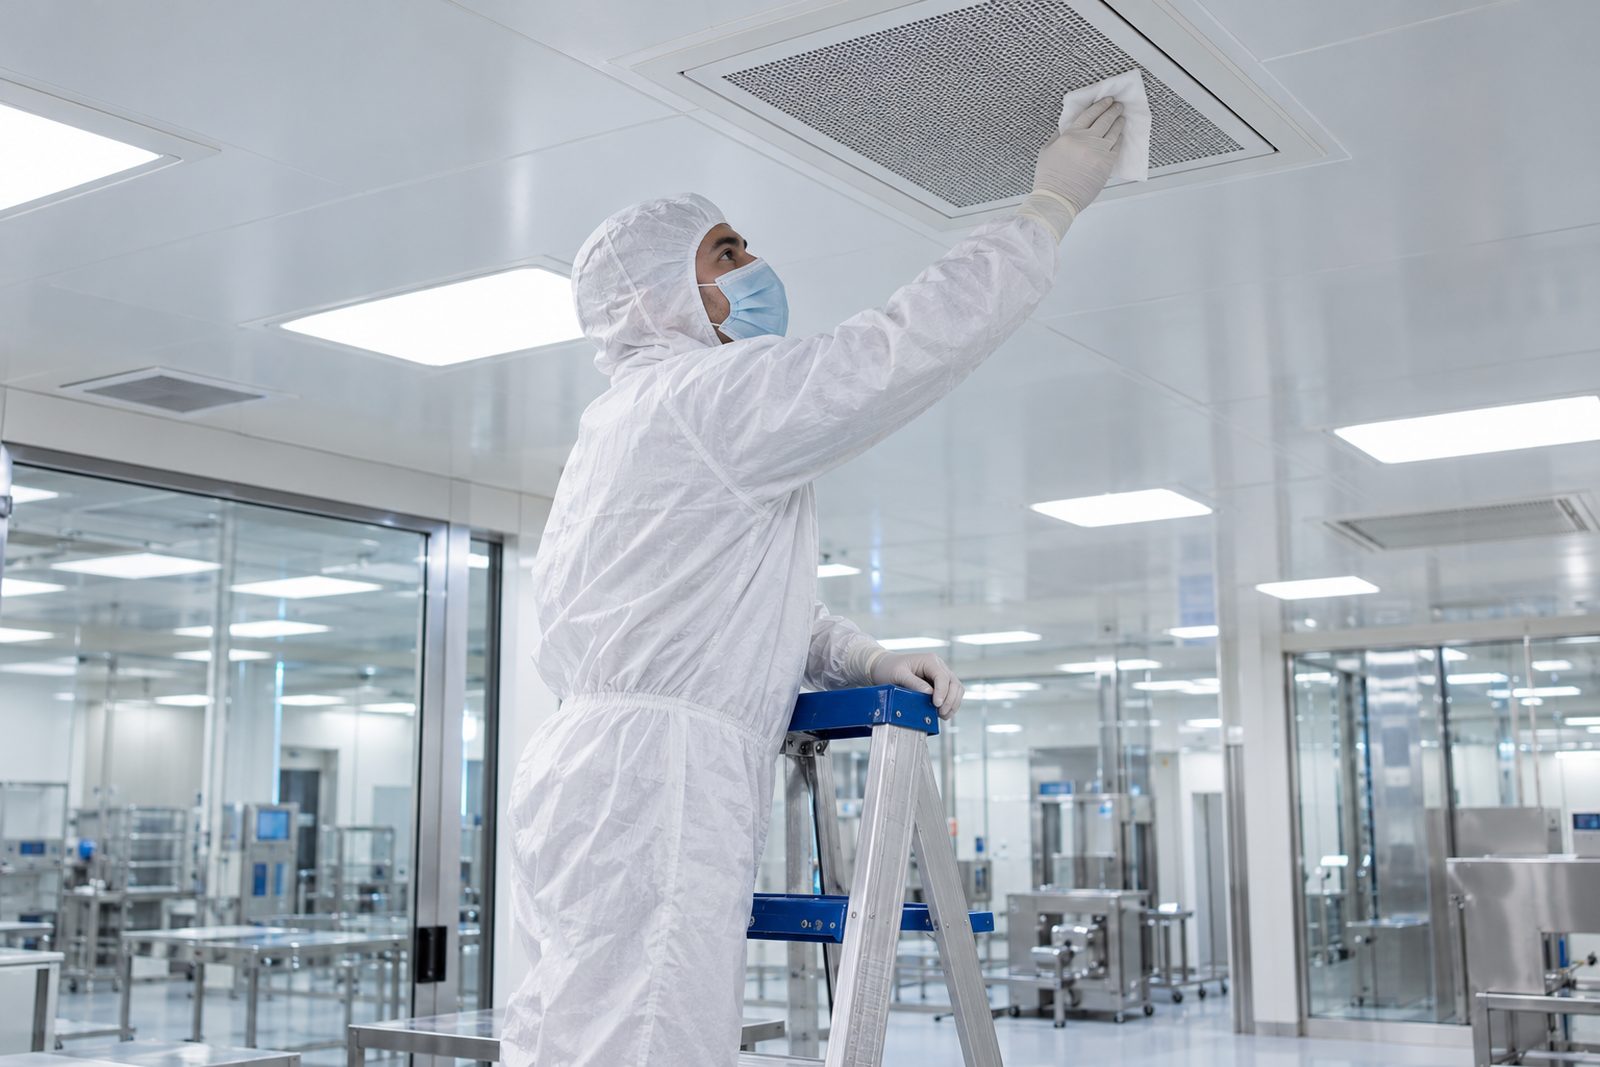
For sterile medicinal-product manufacture, EU GMP Annex 1 describes a written cleaning and disinfection program. When reviewing your program, assess:
- Disinfectant selection and periodic sporicidal use supported by efficacy assessments and monitoring
- Defined surface coverage, including boundaries and areas around equipment
- Tool segregation or decontamination between zones under an approved procedure
- Cleaning logs that align with your EM program so QA can correlate events to cleaning records
- Technicians who understand why procedures exist, not just what steps to follow
Define training, supervision and documentation responsibilities with your cleanroom services provider. Assess contamination risks alongside the rest of the facility quality system.